

Regulation of Medical Devices in Ukraine
In Ukraine, requirements for medical devices are established by technical regulations. They define the essential requirements for product safety and performance, classification rules, conformity assessment procedures, requirements for labelling, technical documentation, and placing devices on the market.
Currently, the regulatory framework is based on three separate technical regulations, depending on the product category:
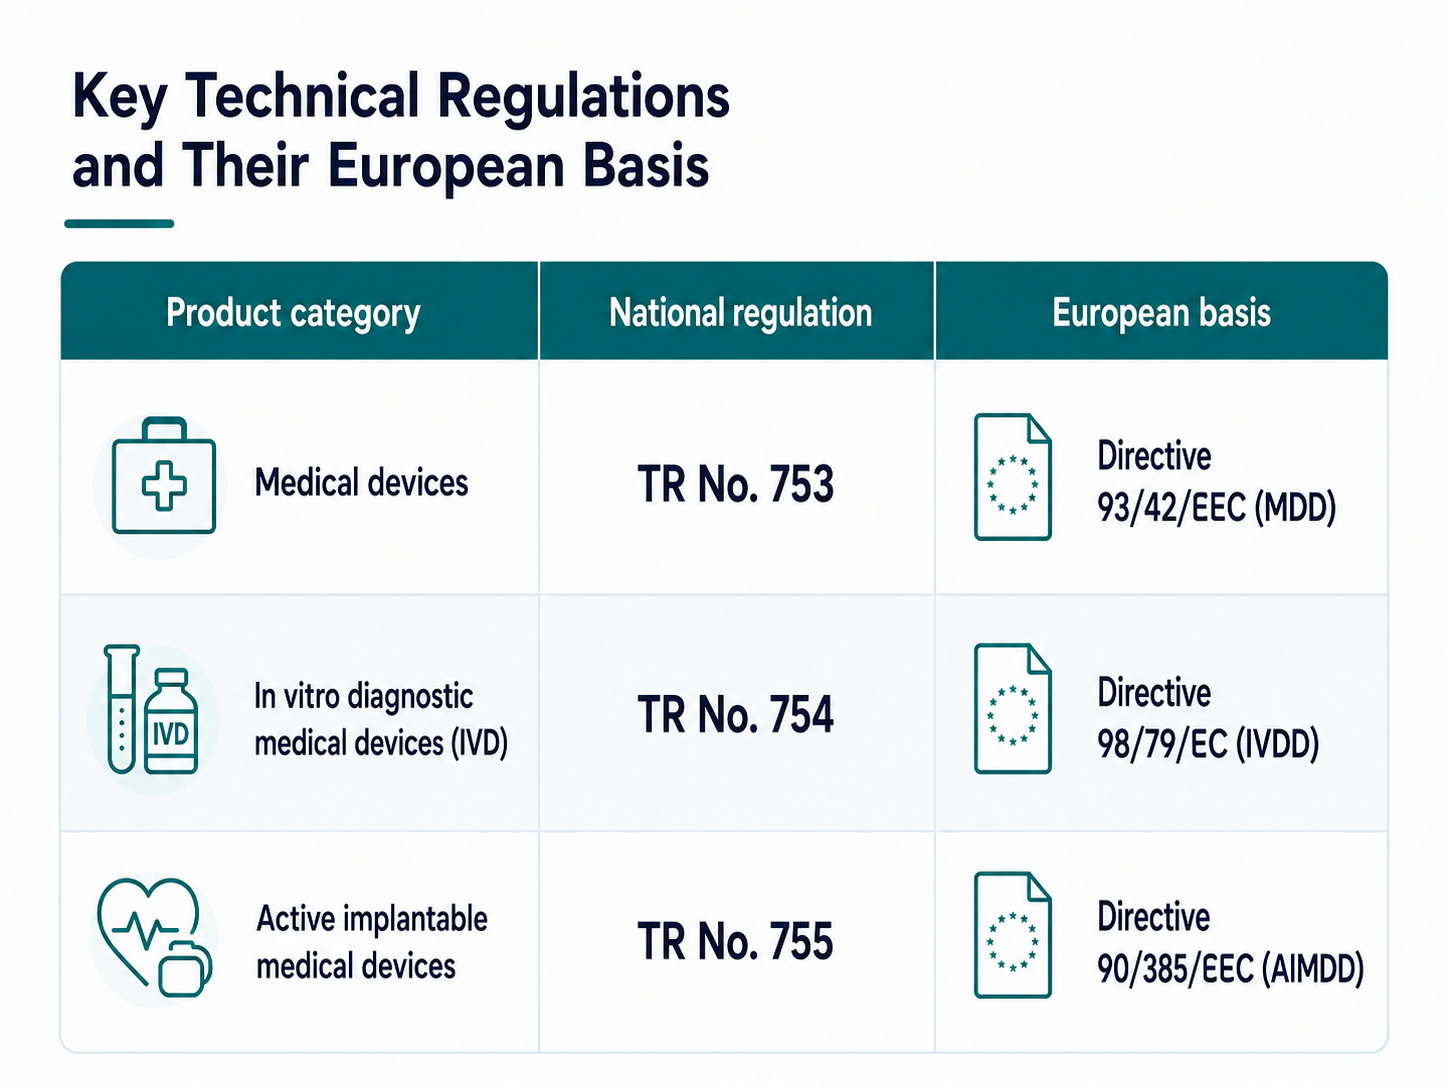
This structure of Ukrainian regulation has brought the requirements for medical devices closer to the European model.
This has practical significance for foreign manufacturers: the structure of technical documentation, the approach to device classification, conformity assessment, and quality management systems are largely based on familiar European regulatory principles. At the same time, the availability of a European EC conformity certificate and the affixing of the European “CE” conformity marking to a medical device do not replace compliance with Ukrainian requirements — before a product is placed on the market, it must undergo the procedure prescribed by Ukrainian legislation.
At the same time, new Technical Regulations will enter into force in December 2026: TR No. 759 on medical devices and TR No. 758 on in vitro diagnostic medical devices.
To allow market participants to prepare for the new rules, a transitional period has been introduced, under which medical devices placed on the market no later than 30 December 2030 may not be prohibited or restricted on the grounds of non-compliance with the requirements of the new technical regulation.
Therefore, until the transition to the new system, conformity assessment of medical devices is carried out under the currently applicable TR No. 753, TR No. 754, and TR No. 755, taking into account the established transitional provisions.
What Is a Medical Device
A medical device is a product used in healthcare for the diagnosis, prevention, monitoring, or treatment of diseases and injuries. It may be an instrument, apparatus, implant, material, or even software. The main difference between a medical device and a medicinal product is that the principal intended action of a medical device is not achieved through pharmacological, immunological, or metabolic effects on the human body.
Medical devices may be placed on the Ukrainian market only after their conformity with the requirements of the applicable Technical Regulation has been assessed.
Main Conformity Assessment Routes for Medical Devices
Depending on the risk class of the medical device, its characteristics, manufacturing process, and available documentation, different conformity assessment procedures may apply. In practice, four main routes can be distinguished.
1. Conformity Assessment through a Manufacturing Audit
This route involves a designated conformity assessment body, which audits the manufacturer’s quality management system and reviews the technical documentation for the medical devices. The audit is conducted at the manufacturer’s premises and, in some cases, at manufacturing sites.
This approach is primarily used for serial production of medical devices. Upon successful completion of the procedure, a conformity assessment certificate is issued for a period of 5 years.During its validity period, annual surveillance audits are conducted, which are considerably simpler and less expensive than the initial certification audit.
2. Conformity Assessment through Recognition of a European Certificate
If a medical device has already undergone conformity assessment in the European Union with the involvement of a European Notified Body, the results of that assessment may be used by a Ukrainian conformity assessment body.
In this case, the Ukrainian conformity assessment body reviews the existing European certificate and the technical file for the medical device. This makes it possible to avoid repeating in full the assessment activities whose results may be recognised in Ukraine and generally makes the procedure simpler, faster, and less expensive compared with a full conformity assessment.
The validity of the Ukrainian conformity certificate is limited by the validity period of the European certificate and by the list of products covered by that certificate.
No manufacturing audit is conducted under this procedure.
3. Self-Declaration and Registration
For medical devices where the legislation does not require the involvement of a designated conformity assessment body, the manufacturer independently confirms that the product complies with the applicable requirements. This approach applies, in particular, to Class I medical devices.
At the same time, self-declaration is not limited to signing a declaration of conformity. Before drawing up the declaration, the manufacturer must assess compliance with the requirements of the Technical Regulation, identify the applicable Essential Requirements, prepare the technical documentation, and ensure that sufficient evidence is available to demonstrate the safety and conformity of the device. The labelling and instructions for use, where required, must also be properly prepared.
A key feature of self-declaration is that a designated body does not review the technical documentation or verify the correctness of the manufacturer’s conformity assessment in advance. Therefore, the manufacturer is responsible for the justification of the declaration and for maintaining documentary evidence of conformity. The relevant documentation must be retained and made available to the market surveillance authority in the event of an inspection.
For Class I medical devices, as well as custom-made medical devices, systems and procedure packs, registration in the State Service of Ukraine on Medicines and Drugs Control Register is required in the cases provided for by law.
4. Conformity Assessment of a Batch of Medical Devices
This route allows conformity of a specific batch of medical devices to be confirmed without conducting a full audit of the manufacturer’s quality management system.
The conformity assessment body examines the products and the necessary documentation relating to the specific batch. The result of such an assessment applies only to the products included in the assessed batch and does not cover the manufacturer’s entire serial production.
This procedure may be advantageous for homogeneous products that are imported infrequently. Where a broad range of products is imported, each individual product item must be assessed, which affects the final cost of the project.
Upon successful completion of the procedure, the conformity assessment body issues a certificate for the batch.
Medical devices that have successfully undergone the conformity assessment procedure under the applicable Technical Regulation must bear the conformity mark to technical regulations.
Classification of Medical Devices
Medical devices are classified into Classes I, IIa, IIb, and III. Classification is carried out in accordance with the criteria set out in Annex 2 to Technical Regulation No. 753. додатку 2 Технічного регламенту № 753
The applicable conformity assessment procedure depends on the class of the medical device.
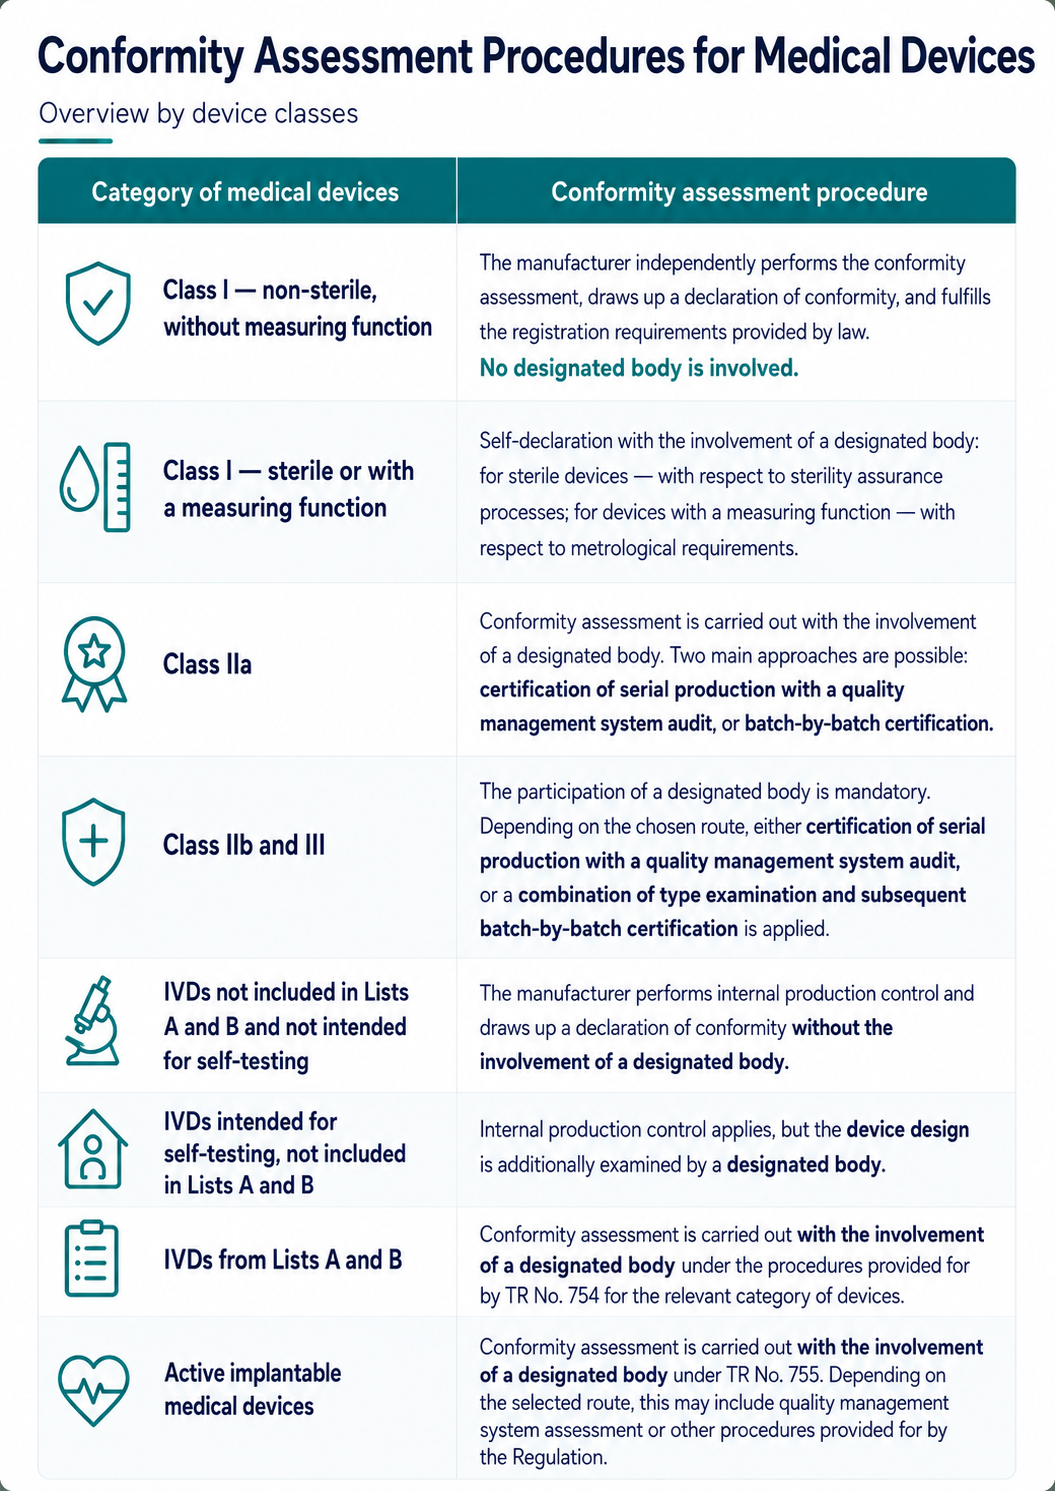
By Order of the Ministry of Health of Ukraine No. 142 dated 22 January 2020, the Methodological Recommendations on the Application of Medical Device Classification Rules were approved.
Designated Conformity Assessment Bodies for Medical Devices
If the conformity assessment procedure requires the involvement of an independent third party, it is carried out by a designated conformity assessment body.In the European Union, similar organizations are referred to as Notified Bodies..
A conformity assessment under the relevant Technical Regulation may be performed by a body that has the necessary accreditation and a valid designation by the Ministry of Economy of Ukraine to perform such activities. The designation determines the specific technical regulations and procedures under which the body is authorized to carry out conformity assessment.
|
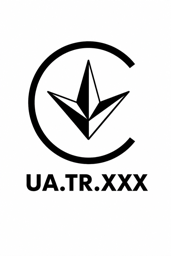
Each designated body is assigned its own identification number in the format UA.TR.XXX.Where a designated body has been involved in the conformity assessment and the Technical Regulation requires its identification number to be indicated, the number is placed next to the conformity mark to technical regulations — on the medical device or its packaging (label), as well as in the instructions for use, where such instructions are mandatory. |
 |
The manufacturer or its Authorized Representative independently selects the designated body that will carryout the conformity assessment.
Although all designated bodies are subject to the same regulatory requirements, in practice their approaches may differ significantly. When selecting a body, it is advisable to consider its experience with specific types of medical devices, requirements for technical documentation, auditor availability, the possibility of conducting audits in a foreign language, timelines, and the cost of services.
For certification of serial production, it is advisable to compare not only the initial cost of obtaining the certificate but also the costs of maintaining it:surveillance audits, assessment of changes, expansion of the product scope or manufacturing sites, and other related activities. Therefore, the choice of a designated body can significantly affect the overall cost, timelines, and convenience of the entire conformity assessment process.
Authorized Representative of a Medical Device Manufacturer
If the manufacturer of a medical device is not a resident of Ukraine, it must appoint an Authorized Representativein order to place its products on the Ukrainian market. The Authorized Representative may be a legal entity or an individual registered in Ukraine to whom the manufacturer delegates certain powers related to compliance with Ukrainian medical device legislation.
The name and address of the Authorized Representative must be indicated on the labelling and in the instructions for use of the medical device. і в інструкції із застосування медичного виробу.
The representative’s authority must be established in writing. In practice, it is advisable to enter into an agreement with the manufacturer defining the functions, rights, and obligations of the parties, as well as to issue a power of attorney. The power of attorney may be used to confirm the representative’s authority before conformity assessment bodies, customs authorities, public authorities, and other third parties.
The Authorized Representative ensures communication between the foreign manufacturer and Ukrainian public authorities, conformity assessment bodies, and other market participants. It also performs the regulatory functions prescribed by law after the product has been placed on the market, including the provision of required documentation and interaction with market surveillance authorities.
For one medical device, the manufacturer appoints one Authorized Representative. At the same time, the legislation does not require a single representative to cover the manufacturer’s entire product portfolio: different Authorized Representatives may be appointed for different medical devices..
The functions of the Authorized Representative may be performed by a Ukrainian importer or distributor of the products. The manufacturer may also engage an independent specialized companythat is not involved in the supply or sale of medical devices and performs exclusively regulatory functions.
Pharma Navigation provides Authorized Representative services to foreign manufacturers of medical devices in Ukraine. This model is particularly convenient for manufacturers working simultaneously with several importers or distributors and wishing to entrust the Authorized Representative function to an independent company.
This allows the manufacturer to maintain a consistent regulatory model in the Ukrainian market regardless of changes in or the number of distributors, while also maintaining better control over access to confidential technical documentation. Engaging an independent Authorized Representative makes it possible to centralize the storage and exchange of such documentation and to disclose it only to the extent necessary to meet regulatory requirements.
Pharma Navigation performs the functions of the Authorized Representative, communicates with the manufacturer and the market surveillance authority, manages regulatory documentation, and provides support on matters related to the placing on the market and subsequent circulation of medical devices in Ukraine.
Labelling and Instructions for Use Requirements
To place a medical device on the Ukrainian market, its labelling and instructions for use must be adapted to the requirements of the Technical Regulation. For imported products, this usually means not merely translating the original packaging, but also verifying whether it contains all information required in Ukraine.
Information about the medical device intended for the consumer or user must be provided in the Ukrainian language.At the same time, other languages may remain on the packaging — for example, the manufacturer’s original multilingual labelling.
The labelling must contain the information necessary to identify the device and ensure its safe use. Depending on the specific device, this may include, among other things, the name and address of the manufacturer, package contents, batch or serial number, expiry date, information on sterility, storage conditions, special conditions of use, warnings, and precautions. If the intended purpose of the device is not obvious to the user, it must also be clearly stated.
For products manufactured outside Ukraine, particular attention should be paid to the details of the Authorized Representative in Ukraine. Technical Regulation No. 753 requires the name and address of the Authorized Representative to be indicated for imported devices. If the medical device is accompanied by instructions for use, the Authorized Representative’s details must also be included in the instructions.
After completing the applicable conformity assessment procedure, medical devices must bear the conformity mark to technical regulations.It is affixed to the medical device or its packaging, as well as to the instructions for use. If a designated body was involved in the conformity assessment, its identification number is indicated next to the conformity mark.
As a general rule, instructions for use must be provided in the packaging of each medical device. However, for Class I and Class IIa devices, TR No. 753 provides an exception: instructions for use may be omitted if the device can be used safely without them.
The content of the instructions depends on the intended purpose, design, and method of use of the particular device. Clause 47 contains a broad list of information to be included where necessary.
There are also two formal requirements that should be checked separately when adapting the instructions. The instructions must include the details of the manufacturer’s Authorized Representative in Ukraine, as well as the date of issue or the date of the latest revision of the instructions.
Therefore, preparing Ukrainian instructions for use is not merely a matter of translating the original document. After translation, the document should be separately reviewed for compliance with TR No. 753 and supplemented with any local information that was not included in the manufacturer’s original documentation.
Symbols Used on Labelling
A significant amount of information on medical devices may be communicated not through text, but by means of standardized graphical symbols. For this purpose, DSTU EN ISO 15223-1:2022 “Medical devices. Symbols to be used with information to be supplied by the manufacturer. Part 1: General requirements” is applied. The standard has been effective in Ukraine since 31 December 2023 and replaced the previous edition, DSTU EN ISO 15223-1:2018.
Such symbols include, for example, those indicating the manufacturer, date of manufacture, batch number, serial number, expiry date, sterility, single use, and the need to consult the instructions for use. The use of standardized symbols allows packaging to be designed more compactly while ensuring that key information remains understandable to users in different countries.
Advertising and Promotion of Medical Devices
Advertising of medical devices in Ukraine is permitted provided that the relevant device is authorized for use in Ukraine.The main specific requirements are established by the Law of Ukraine “On Advertising”, while the accuracy of advertising claims should also be assessed with due regard to legislation on protection against unfair competition.
Unlike medicinal products, medical devices are not divided into prescription and non-prescription categories for the purpose of determining whether they may be advertised. At the same time, advertising must contain objective information about the device and must not exaggerate its properties.In particular, claims of a guaranteed therapeutic effect, positioning a device as the “most effective” or “safest”, references to specific successful cases of use, recommendations by healthcare professionals, as well as information capable of misleading consumers as to the characteristics or effectiveness of the device, are prohibited. Advertising must also not feature physicians or persons whose appearance imitates that of physicians.
The permissibility of advertising claims is assessed by the advertiser itself before publication.
Advertising intended for the general public should be distinguished from professional promotion addressed to medical and pharmaceutical professionals. For materials published in specialized professional publications or on specialized websites, the Law “On Advertising” provides for a separate regime, and some of the general restrictions set out in Article 21 do not apply to such communications.
Before publishing advertising and promotional materials, it is advisable to verify not only their formal compliance with the law, but also the evidentiary basis for each claimed effect. Pharma Navigation performs regulatory reviews of advertising materials, assesses the permissibility of claimed properties, and helps adapt marketing claims to the requirements of Ukrainian legislation.
Application of Other Technical Regulations
In certain cases, a medical device may simultaneously fall within the scope not only of the sector-specific Technical Regulation on medical devices, but also of other technical regulations. In such cases, the manufacturer must take into account account all requirements applicable to the specific deviceduring conformity assessment and preparation of the technical documentation. The legislation allows a declaration of conformity to refer to several technical regulations where more than one regulation applies to the product.
For medical devices, the following regulations may be particularly relevant:
- Technical Regulation on the Restriction of the Use of Certain Hazardous Substances in Electrical and Electronic Equipment (RoHS), approved by Resolution of the Cabinet of Ministers of Ukraine No. 139. It applies, in particular, to electrical and electronic medical devices, including IVDs. At the same time, the Regulation contains its own exclusions — in particular, active implantable medical devices are excluded from its scope.
- Technical Regulation on Radio Equipment (RED), approved by Resolution of the Cabinet of Ministers of Ukraine No. 355. It should be taken into account where a medical device has radio communication functions and falls within the definition of radio equipment — for example, where it uses Bluetooth, Wi-Fi, or other wireless data transmission technologies.
Therefore, when determining the conformity assessment route, it is important to assess not only the medical purpose of the device, but also its technical characteristics. The same product may simultaneously be subject to the requirements of the medical device Technical Regulation, RoHS, and the Technical Regulation on Radio Equipment.
Particular attention should also be paid to medical devices with a measuring function. Technical Regulation No. 94 on Legally Regulated Measuring Instruments does not apply to measuring instruments that are medical devices or in vitro diagnostic medical devices. Requirements relating to their measuring function are assessed within the framework of the applicable medical device regulation.
Market Surveillance
Once a product has been placed on the market, it becomes subject to state market surveillance,during which its actual compliance with the requirements of the technical regulations may be verified.
Until 31 December 2026, the State Service of Ukraine on Medicines and Drugs Control remains the state market surveillance authority for medical devices. From 1 January 2027, these functions will be transferred to the Ukrainian Pharmaceutical Agency (UPA), which has been established as the legal successor to the State Service and, among other functions, will be responsible for technical regulation and market surveillance of medical devices.
Market surveillance may be carried out through planned inspections in accordance with the sectoral plan, as well as through unscheduled measures in cases provided for by law. Inspections are carried out directly at the premises of participants in the supply chain — including distributors of medical devices. Separately, when products are imported into Ukraine, customs authorities carry out state control of products and may interact with the market surveillance authority.
In practice, particular attention is paid to labelling, instructions for use, and documents confirming the conformity of the device. Typical violations identified by the State Service include the absence of Ukrainian-language information on the packaging, the conformity mark, Authorized Representative details, information necessary to identify the device, instructions for use, or the date of issue or latest revision of the instructions.
Identification of a violation is not limited to the imposition of a fine. Depending on the nature of the violation and the level of risk, the market surveillance authority may require the economic operator to correct the non-compliance, restrict or prohibit the making available of the product on the market, withdraw it from the market, or recall products that have already been supplied.
The amount of the fine depends not only on the seriousness of the violation, but also on the type of violation, the status of the economic operator, and whether the violation is repeated. Fines are imposed for each model, article, or batch of products that is dangerous, poses a risk and/or does not comply with the established requirements, regardless of the number of units of such products and/or the number of places where they are sold.
Our Services
Pharma Navigation supports manufacturers and importers of medical devices at all stages of conformity assessment and preparation of products for placing on the Ukrainian market. We help determine the applicable requirements and the optimal conformity assessment route, organize interaction with the designated body, analyze the available documentation, and coordinate the preparation of documents required for the procedure.
Within the project, we may support communication between the manufacturer, the Authorized Representative, and the conformity assessment body, assist with the preparation of the declaration of conformity, adaptation of labelling and instructions for use, as well as other regulatory matters arising when placing a medical device on the Ukrainian market.
In addition, Pharma Navigation provides:
- Authorized Representative services for foreign manufacturers in Ukraine;
- legal and regulatory support for manufacturers, Authorized Representatives, importers, and distributors of medical devices.
This approach allows the coordination of the regulatory process to be entrusted to a single team and minimizes the need for independent interaction with different participants in the conformity assessment procedure.